Short answer Medical device registration in India means finding your device’s risk class and licence route under the Medical Devices Rules, 2017, preparing technical and quality documents, submitting the application on CDSCO’s Medical Devices Online portal, answering regulatory queries, and receiving the applicable licence or registration number. The route differs for importers, manufacturers and Class A to D devices.
Many guides describe this as one fixed process. It is not. Your path depends on the device’s class, its intended use, and whether you manufacture in India or import. The seven steps below follow the order in which the rules work, so you can see where your own product fits.
Registration, licence, approval: what each word means
- Registration
- For Class A non-sterile, non-measuring devices, a system-generated registration number from the portal. People also use the word loosely for the whole process. Older “registration numbers” issued for notified devices are no longer treated as approval, says CDSCO.
Licence- The legal permission to import or manufacture most devices, such as Form MD-15 (import) or MD-5 and MD-9 (manufacture).
Approval- A wider word. It also covers permissions such as clinical investigation (Form MD-23).
- “CDSCO registration”
- An everyday phrase. CDSCO is the national regulator, but Class A and B manufacturing licences are issued by State Licensing Authorities.
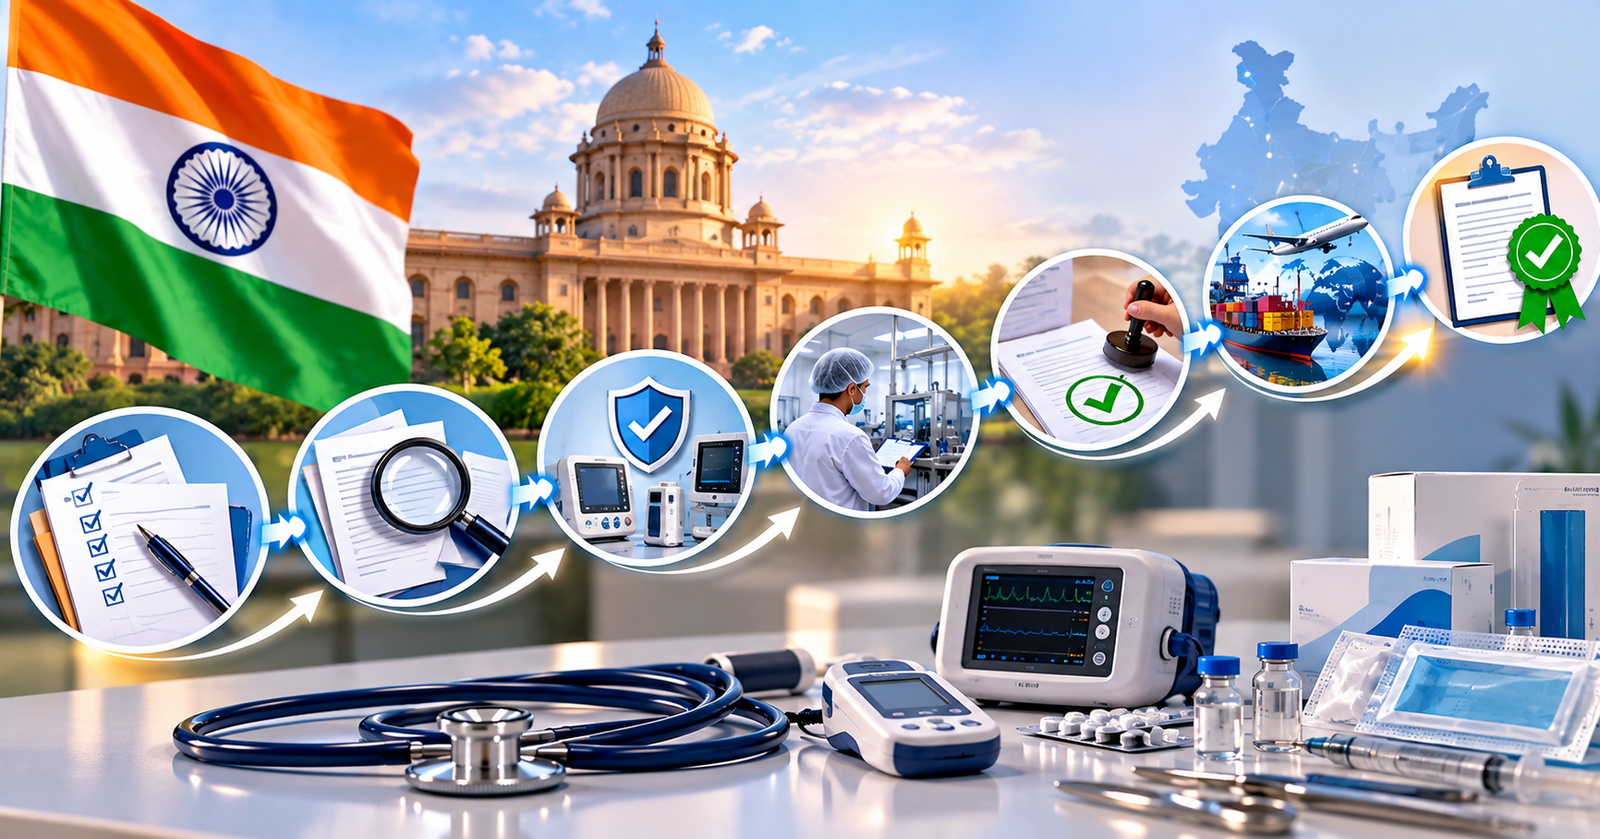
The seven steps at a glance.
1
Confirm your product is a medical device
All medical devices in India are regulated under the Medical Devices Rules, 2017, following notification S.O. 648(E) of 11 February 2020. So the first question is whether your product meets the definition. The manufacturer’s stated intended use decides this. Standalone software can qualify too.
Why it matters. A wellness massager is outside the rules, says CDSCO, but one meant to treat a condition is inside them. Your claims on labels and marketing material can change the answer.
Common mistake. Copying claims from a competitor’s brochure. Write your own intended-use statement and keep it identical everywhere.
2
Confirm the risk class: A, B, C or D
Devices are classified by risk: Class A is low, B low-moderate, C moderate-high, D high. Only the Central Licensing Authority (the Drugs Controller General of India at CDSCO) classifies devices, using the First Schedule and a published list that is updated from time to time.
Why classification comes first. It decides the licensing authority, the forms, the documents, whether a site audit happens and the fee. Guessing wrong sends every later step in the wrong direction.
If your device is not on the list, CDSCO says you can send a request with the label, instructions for use and the product’s regulatory status in other countries. For imports, the higher class may apply when India and IMDRF countries classify differently.
Tip. Download the classification list on the day you start, not from an old folder. Lists are revised, so an old copy can mislead you.
Identify your licence route
- Now combine two facts: what you are (importer or manufacturer) and your class. That gives you the form and authority.Overseas manufacturers cannot simply apply themselves. Import goes through an authorised agent in India, appointed by power of attorney, who must hold a manufacturing, wholesale or MD-42 licence. Selling and distribution have their own licence or registration under the State Licensing Authority.
Your situation Application → Licence Authority Import, any class MD-14 → MD-15 Central (CDSCO) Manufacture Class A (sterile or measuring) or B MD-3 → MD-5 State Licensing Authority Manufacture Class C or D MD-7 → MD-9 Central (CDSCO) Class A non-sterile, non-measuring Portal registration, no licence, no fee Medical Devices Online portal Prototype or test batches MD-12 → MD-13 (manufacture); MD-16 → MD-17 (import) Central (CDSCO) New device with no predicate Clinical investigation permission MD-22 → MD-23; then MD-26 → MD-27 Central (CDSCO) Common mistake. Assuming Class A and B are filed with CDSCO. Manufacturing of these goes to the state where your factory is located.
4
Prepare the regulatory and technical documents
The portal has a checklist for each form, and CDSCO says that checklist is the reference for essential documents. The content comes from the Fourth Schedule of the Rules. Exact documents depend on your route and class, but these commonly appear:
- Device Master File (DMF): design, specifications, risk management, performance data, labels and instructions for use.
- Plant Master File (PMF) or Site Master File: your manufacturing site details (manufacturers).
- Essential Principles checklist: safety and performance principles apply to every risk class.
- For import: power of attorney with the agent’s undertaking, authenticated as the Rules require, and a Free Sale Certificate naming both the legal and actual manufacturer. A Certificate of Exportability is not accepted in its place.
- Labels and instructions for use meeting Chapter VI of the Rules.
Common mistake. Small mismatches: a product name, address or model number that differs between the DMF, the certificate and the label. Reviewers notice these.
Tip. Ask for a separate fee check per brand. CDSCO states that each brand of a device needs its own fee.
5
Complete testing, quality system and inspection readiness
Testing. Send devices for test to a Medical Device Testing Laboratory registered with CDSCO. Where a BIS standard exists for the device, meeting it is mandatory. If none exists, applicable ISO or IEC standards apply. Devices with a claimed shelf life need real-time ageing data, or accelerated data with real-time studies under way.
Quality system. Manufacturers must comply with the Quality Management System requirements in the Fifth Schedule and submit a signed undertaking. Importers submit a quality certificate as the Fourth Schedule requires. ISO 13485 aligns closely with these requirements, but check whether your route asks for a certificate or only compliance.
Audits. This differs by class. No pre-licence site audit applies to Class A manufacturing. For Class B, a notified body assigned by the State Licensing Authority audits the site. For Class C and D, CDSCO’s Medical Device Officers inspect. CDSCO may also inspect an overseas site before or after an import licence.
Clinical evidence. A clinical investigation is needed only for investigational devices, meaning those without a predicate, or licensed devices with a new intended use or major change.
6
Submit online, pay fees and answer regulatory queries
Applications go through CDSCO’s Medical Devices Online portal, also reachable through the National Single Window System. Fees are set in the Second Schedule of the Rules, and CDSCO says paid fees are not refunded if you withdraw. Only an authorised person of the company can ask for the status of an application, so the company stays the applicant even when a consultant prepares the file.
After filing, officers review the file. They may raise queries or, for some routes, audit observations. Answer each point directly, attach evidence, and keep documents consistent with what you filed.
Common mistake. Answering a query with a new document that contradicts the original dossier. Fix the source document, then reply.
On timing. The Rules give an outer period of nine months for import licence grant, and CDSCO FAQs say Class C and D inspections happen within 60 days when data is satisfactory. Real duration varies with class, completeness, audits and how fast you reply.
7
Receive your licence and stay compliant
When the authority is satisfied, you receive the licence or registration number for your route. Licences are not renewed every five years. Under CDSCO’s FAQ, they stay valid until suspended or cancelled, but the holder must pay a retention fee every five years.
After grant, your duties continue:
- Report major changes for approval, and inform CDSCO of minor changes within 30 days.
- Apply for an endorsement to add products to an existing licence.
- Submit post-marketing surveillance data, as the licence holder.
- Keep labels compliant, and pay retention fees on time.
- Inform the authority within 45 days of a change in company constitution.
Tip. Keep a calendar for retention dates and change triggers. Missing one can weaken an otherwise sound licence.
Common reasons medical device registration gets delayed
- Wrong class or wrong licence route selected at the start.
- Names, addresses or models that differ between documents.
- Power of attorney or Free Sale Certificate that does not meet the stated requirements.
- Test reports that do not come from an accepted laboratory or standard.
- Quality system gaps found at audit and not closed on time.
- Slow or incomplete replies to authority queries.
How a regulatory consultant can help
A consultant does not grant approval. CDSCO and the State Licensing Authorities do. A consultant can help you check classification, choose the route, build the dossier, prepare for audits, draft query replies and plan post-licence compliance. Outcomes and timelines are never guaranteed by anyone outside the regulator.
Read Also: CDSCO Compliance Challenges in India
How TRACC Global can support you
TRACC Global Research and Consulting Pvt Ltd is a regulatory consultancy based in Sector 45, Gurugram. Its website lists support for CDSCO import and manufacturing licence applications, technical documentation, quality management system setup aligned with ISO 13485, portal filing support, and coordination with the Indian authorised agent for overseas manufacturers. You can read more on its pages for CDSCO medical device registration, import licence, manufacturing licence and ISO standards.
Need assistance with medical device registration and regulatory requirements in India? Connect with TRACC Global through https://traccglobal.com/
Frequently Asked Questions
Does every medical device need a licence?
How much does registration cost?
How long does it take?
What if my device is not on CDSCO’s classification list?
Can a foreign manufacturer apply directly?
Company credentials
ISO 9001, ISO 27001 & ISO 45001 certified consultancy
Contact
info@traccglobal.com